Epigenetics data analysis: methylation, chromatin and 3D genome
DNA methylation, chromatin accessibility, transcription factor binding and 3D genome organization: Sycomia analyzes your epigenomics data to reveal the regulation behind your biological question.
DNA methylation
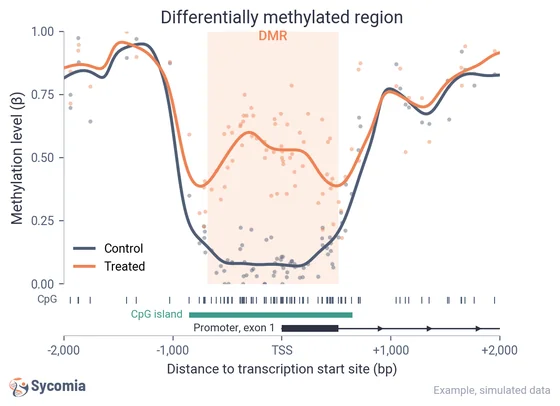
DNA methylation is a chemical mark on DNA that modulates gene activity without changing the sequence. It varies with tissue, age, environment and disease, which makes it a source of biomarkers.
Example analyses
- WGBS, RRBS and EM-seqHigh-quality read alignment and methylation calling for whole-genome bisulfite, reduced representation bisulfite and enzymatic (EM-seq) data.
- Differential methylationRobust identification of regions with significant methylation changes between conditions, for epigenetic biomarker discovery.
- Epigenetic clocksEstimation of biological age with methylation clocks, and of age acceleration and its links with health, disease and exposures.
- Long-read methylationMethylation detected directly on Oxford Nanopore and PacBio long reads, without chemical conversion, resolved by allele and by haplotype.
Chromatin accessibility and TF binding
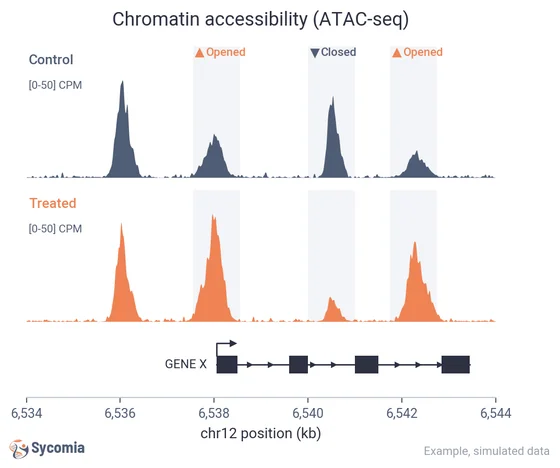
DNA is packaged into chromatin, which can be more or less compact. ATAC-seq finds open regions, accessible to transcription factors; ChIP-seq and CUT&Tag locate a given protein or histone mark across the genome.
Example analyses
- ATAC-seq, ChIP-seq, CUT&TagAssay-specific quality control: fragment size distribution, TSS enrichment, fraction of reads in peaks (FRiP) and mitochondrial reads for ATAC-seq; deduplication and blacklist filtering for every assay.
- Peak callingPrecise detection of enriched regions to reveal regulatory elements, transcription factor binding sites and histone modifications.
- Motifs and footprintingEnriched DNA sequence motifs, transcription factor footprints and motif-level accessibility changes, to identify the transcription factors at work.
- Differential accessibility and bindingRegions that open, close or gain binding between your conditions, tested on a consensus peak set with statistical models suited to count data.
3D genomics
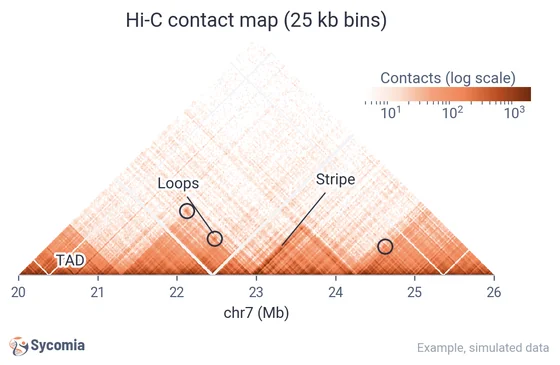
In the nucleus, the genome folds in three dimensions: regions far apart on the DNA can touch and regulate a gene at a distance. Hi-C measures these contacts and reveals compartments, domains (TADs) and chromatin loops.
Example analyses
- Hi-C and Capture Hi-CMapping of chromatin interactions genome-wide (Hi-C) or targeted to promoters and regions of interest (Capture Hi-C).
- Compartments, TADs and loopsIdentification of A/B compartments, Topologically Associating Domains and chromatin loops, and their role in gene regulation.
- Differential analysisComparison of compartments, TADs and loops between conditions, with clear contact maps to show the changes.
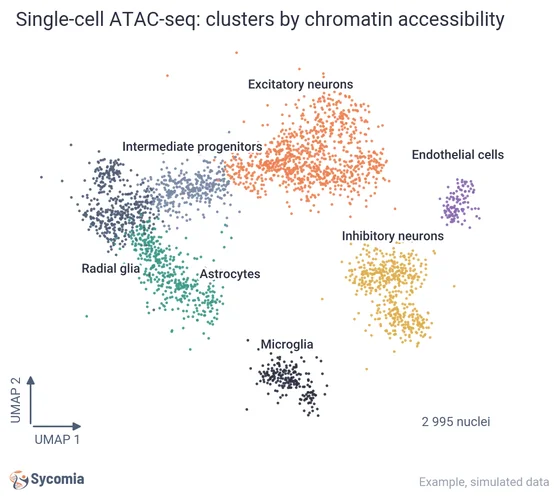
Single-cell epigenomics
Single-cell epigenomics measures chromatin accessibility or methylation cell by cell. It reveals the regulatory programs of each cell type, including in rare populations.
Example analyses
- scATAC-seq and single-cell methylationAnalysis of chromatin accessibility and DNA methylation (scBS-seq, snmC-seq) at single-cell resolution.
- Epigenetic trajectoriesReconstruction of the dynamic changes of chromatin or methylation along differentiation paths.
- Integration with scRNA-seqJoint analysis linking gene expression with chromatin accessibility or methylation in the same populations.
Your analysis is not on the list? We also build custom analyses, tailored to your biological question. Let’s talk
Frequently asked questions
Another question about your project? Write to us, an expert will answer.
Contact usWhich DNA methylation technologies do you analyze?
Whole-genome bisulfite sequencing (WGBS), reduced representation bisulfite sequencing (RRBS), enzymatic methyl sequencing (EM-seq), and direct detection on Oxford Nanopore and PacBio long reads.
Can you estimate biological age from methylation data?
Yes. We use DNA methylation clocks to estimate biological age and assess age acceleration and its associations with health, disease or exposures.
Do you analyze single-cell epigenomics data?
Yes: scATAC-seq and single-cell DNA methylation (scBS-seq, snmC-seq), epigenetic trajectories, and integration with single-cell RNA-seq.
Is my data kept confidential?
Yes. We sign an NDA on request, before any exchange of data. Your data is processed on our own servers, clinical data is hosted in France by a certified health data host (HDS), and your data is deleted within 30 days of the end of the project, or earlier on request, in line with the GDPR.
Related services
Let’s talk about your epigenetics project
Methylation, chromatin or 3D genome: tell us about your biological question and your data, and we will propose the right analysis strategy.